Aligning Protein Structures with GR-Align (GRaphlet-based Aligner)
Noël Malod-Dognin and Nataša Pržulj
Corresponding author: Dr. N. Malod-Dognin, noel.malod@mbzuai.ac.ae
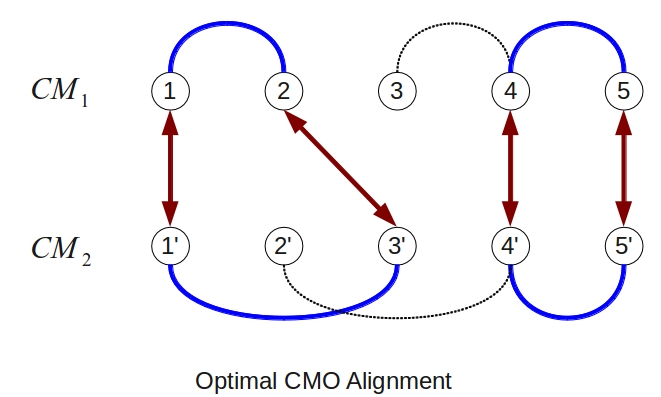
Optimal CMO alignment
GR-Align is a fast heuristic for aligning and comparing protein structures under the Contact Map Overlap maximization (CMO) paradigm. It uses a Needleman-Wunsch algorithm in which the cost of matching two residues is their graphlet degree similarity.
Version 1.5: outputs of the original software were hard to parse. Aligned residues are now labelled with their PDB IDs.
Downloads
How to run GR-Align
The package contains three programs:
-
CMap converts protein structures (
.pdb) into contact map graphs (LEDA graphs,.gw) -
DCount computes graphlet degree signatures (
.ndump) from contact maps - GR-Align aligns protein structures (needs both contact maps and signatures)
Run any of them without parameters (or with -h) to see usage directions.
Example
Create a contact map from chain A of structure 1amk:
./CMap -i 1amk.pdb -c A -o 1amkA.gw -d 7.5
Compute its graphlet degree signature:
./DCount -i 1amkA.gw -o 1amkA.ndump
Align all structures of the Skolnick set, listed in skolnick.lst, with contact maps and signatures in ./skolnick:
./GR-Align -q skolnick.lst -r ./skolnick -o results.txt
Citation
N. Malod-Dognin and N. Pržulj. GR-Align: fast and flexible alignment of protein 3D structures using graphlet degree similarity. Bioinformatics 30(9), 1259–1265 (2014). https://doi.org/10.1093/bioinformatics/btu020