Fuse: Multiple Network Alignment via Data Fusion
Vladimir Gligorijević, Noël Malod-Dognin and Nataša Pržulj
Corresponding author: Prof. N. Pržulj, natasa.przulj@mbzuai.ac.ae
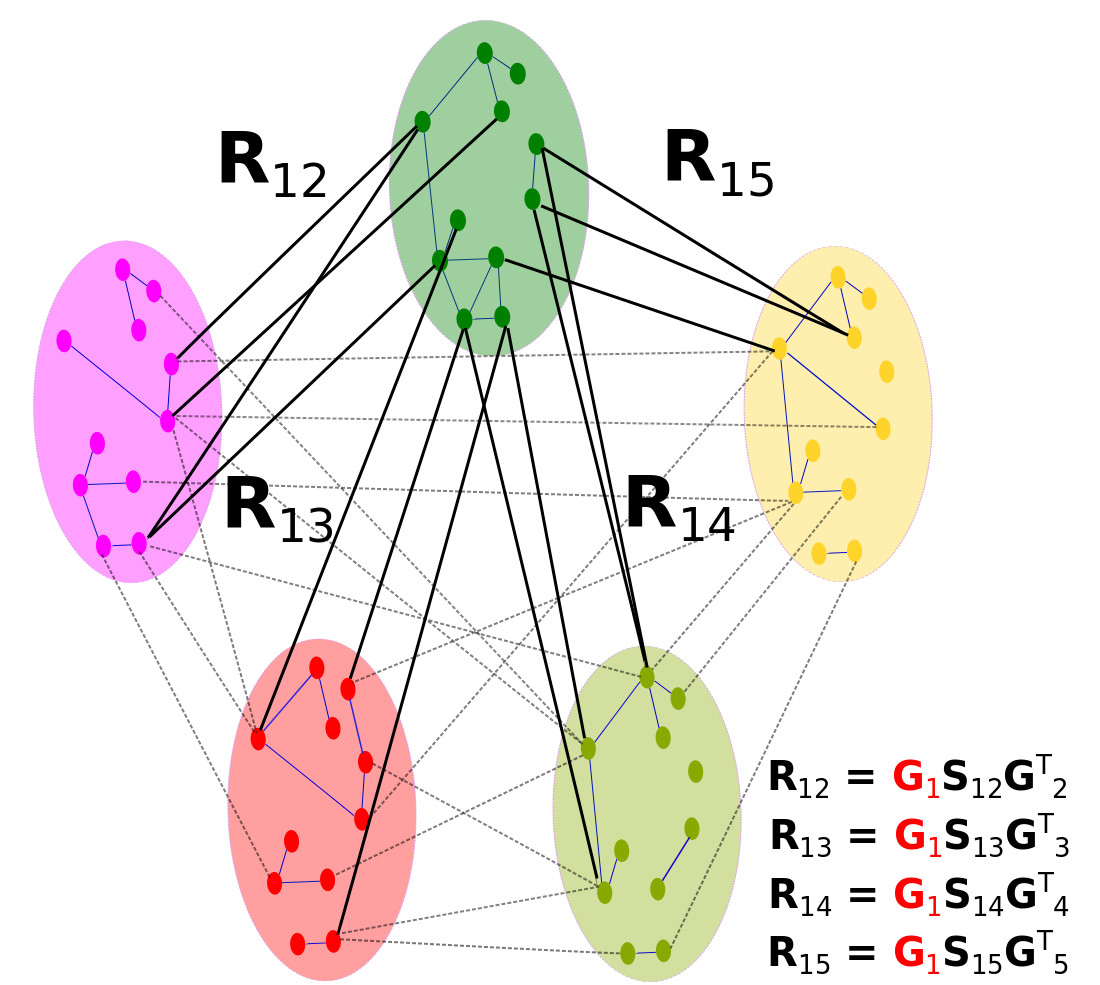
Fuse is a global multiple network aligner. First, it computes new similarity scores between proteins by fusing sequence similarities and network wiring patterns over all proteins in all PPI networks being aligned, using non-negative matrix tri-factorization (NMTF). Second, it constructs a one-to-one global multiple network alignment using an approximate maximum weight k-partite matching solver.
Downloads
How to run Fuse
Step 1: Data fusion (MATLAB)
You need:
- MATLAB R2013a or newer
- the PPI network files, in edgelist format
- a file with the sequence similarities between proteins (e.g. 1 − e-value)
- the number of NMTF iterations, the list of rank parameters, and the gamma parameter
Example (in the MATLAB console):
run_nmtf({'./Nets/CElegans.edgelist', './Nets/DMelanogaster.edgelist', './Nets/HSapiens.edgelist', './Nets/MMusculus.edgelist', './Nets/SCerevisiae.edgelist'}, 'sequence_scores.lst', [80, 90, 80, 70, 50], 1000, 0.01, 'nmtf_scores.lst')
This produces nmtf_scores.lst, the input for the alignment step.
Step 2: Alignment
You need:
- the PPI network files, in LEDA format (
.gw) - a network list file with the path to each network, one per line, e.g.:
./Nets/CElegans.gw ./Nets/DMelanogaster.gw ./Nets/HSapiens.gw ./Nets/MMusculus.gw ./Nets/SCerevisiae.gw - the sequence similarity file and the NMTF-based similarity file from step 1
Example:
./FUSE.exe -n network_list.txt -s sequence_scores.lst -t nmtf_scores.lst -o an_output_file
Run ./FUSE.exe -h for the full list of parameters.
Other multiple network alignment algorithms
Citation
V. Gligorijević, N. Malod-Dognin and N. Pržulj. Fuse: multiple network alignment via data fusion. Bioinformatics 32(8), 1195–1203 (2016). https://doi.org/10.1093/bioinformatics/btv731